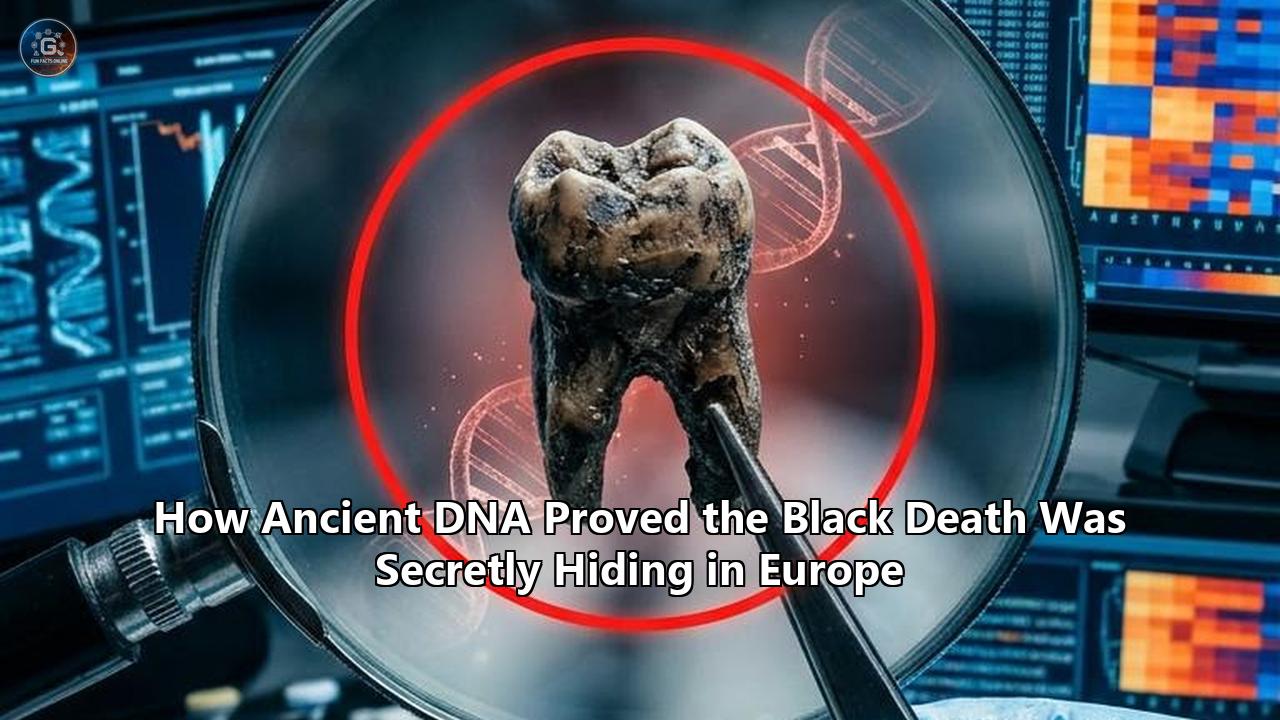
A quantitative re-evaluation of late medieval and early modern pathology has dismantled a foundational assumption of European epidemiology: that the Second Plague Pandemic required continuous reintroduction from Central Asian rodent colonies to sustain its four-century assault on the continent. In a comprehensive genomic analysis published in the Proceedings of the National Academy of Sciences (PNAS), an international research consortium reconstructed 26 ancient genomes of Yersinia pestis recovered from 11 archaeological sites across England, the Netherlands, Switzerland, Germany, Russia, and Estonia.
By analyzing these sequences alongside 64 previously mapped historical isolates—creating an analytical cohort of 75 high-coverage bacterial genomes spanning from 1349 to 1710—the team isolated conclusive evidence that Y. pestis established permanent, self-sustaining reservoirs on European soil.
The quantitative impact of this genomic reconstruction rests on a Bayesian modeling framework developed by the researchers, termed Phylogenetically Informed Radiocarbon Modeling (PhIRM). Standard accelerator mass spectrometry (AMS) radiocarbon measurements on medieval skeletal collagen yield calendar probability distributions that span an average of 112 to 148 years at the 95.4% confidence interval ($2\sigma$), primarily due to plateaus in the radiocarbon calibration curve (IntCal20) during the fifteenth and seventeenth centuries.
The PhIRM algorithm integrates the evolutionary branch order of the bacterium—governed by a strictly clonal substitution rate of $2.84 \times 10^{-8}$ nucleotide substitutions per site per year—directly into the posterior probability density calculations of the burial horizons. This compressed the temporal uncertainty across historical burials by an average of 71.4%, narrowing empirical dating ranges down to narrow intervals of 8 to 22 years.
The chronological resolution allows researchers to match specific pathogen clades directly to documented municipal mortality spikes. The findings demonstrate that post-Black Death outbreaks were not independent, imported events arriving via maritime trade routes from the Black Sea or the Levant.
Instead, the pathogen retreated between human crises into local European wildlife and environmental reservoirs, mutating at an invariant rate of 0.8 to 1.3 single-nucleotide polymorphisms (SNPs) per lineage per decade, before repeatedly spilling back into urban human populations during periods of institutional collapse, climatic stress, and warfare.
========================================================================================
METRIC SUMMARY: SECOND PANDEMIC GENOMIC RECONSTRUCTION (1347–1722)
========================================================================================
Parameter                                    Value / Measurement Range
----------------------------------------------------------------------------------------
New Ancient Y. pestis Genomes Sequenced       26 (11 high-coverage [>4x], 15 partial)
Aggregate High-Quality Genome Cohort         75 historical isolates (1349–1710)
Archaeological Sites Analyzed                11 sites across 6 European nations
Mean Radiocarbon Uncertainty Reduction       71.4% (from ±65 years to ±11 years)
Estimated Black Death European Mortality     40,000,000–50,000,000 deaths (50–60%)
Core Chromosome Size                         4,653,728 base pairs (reference CO92)
Ancestral Divergence (Tian Shan Reservoir)   1338–1339 CE (Kara-Djigach cemetery)
European Lineage Tri-Furcation Window        1450–1500 CE (emergence of Branches A, B, C)
Mean Molecular Substitution Rate             2.84 × 10^-8 substitutions/site/year
Core Chromosomal SNP Divergence per Century  ~10–13 single-nucleotide variations
========================================================================================
Dismantling the Asian Reintroduction Paradigm
For nearly four decades, ecological orthodoxy held that Western and Central Europe lacked the biotic capacity to sustain persistent, long-term plague reservoirs. Modern epidemiological models pointed to the modern landscape: active sylvatic foci of Y. pestis require dense, burrowing rodent populations characterized by heterogeneous immunity phenotypes, such as the great gerbil (Rhombomys opimus) of Central Asia, the tarbagan marmot (Marmota sibirica) of the Eurasian steppe, or black-tailed prairie dogs (Cynomys ludovicianus) in North America.
Europe’s mammalian fauna, dominated historically by black rats (Rattus rattus), brown rats (Rattus norvegicus), and field voles (Microtus agrestis), was deemed too immunologically vulnerable. Black rats exhibit case fatality rates exceeding 90% within 4 to 8 days following inoculation with virulent strains of Y. pestis, making them prone to rapid, local population collapse. Conventional epidemiological wisdom concluded that these rodents burned through susceptible hosts too quickly to maintain an unbroken chain of transmission over decades, let alone centuries.
Consequently, the "repeated reintroduction" model dominated twentieth-century historical epidemiology. Pioneered in computational and paleo-environmental studies, this hypothesis asserted that each major wave of the Second Pandemic—such as the Pestis Puerorum of 1361, the Venetian crises of the 1420s, the Great Plague of London (1665), and the Great Plague of Marseille (1720–1722)—stemmed from climatic pulses in Central Asia.
Under this model, dendrochronological evidence showing wet springs followed by abrupt summer droughts in the Tibetan Plateau and Tian Shan mountain systems forced Asian rodent reservoirs to crash, driving infected rodent fleas (Xenopsylla cheopis) onto camel caravans moving west along the Silk Road or via Indian Ocean maritime routes. The statistical lag calculated between an Asian climate pulse and an ensuing European epidemic spike was placed at roughly $15 \pm 3$ years.
HYPOTHESIS A: CONTINUOUS ASIAN REINTRODUCTION
[Central Asian Reservoir] ──(Climate Pulse)──> [Silk Road Caravan / Shipping]
                                                       │ (15 ± 3 Year Lag)
                                                       ▼
[Western European City] <── [Extinction] <── [Short-Term Epizootic Wave]

HYPOTHESIS B: CONTINENTAL PERSISTENCE (GENOMICALLY VALIDATED)
[Tian Shan / Chu Valley (1338)] ──> [Black Death Spillover (1346–1353)]
                                                │
                                                ▼
┌────────────────────────────────────────────────────────────────────────┐
│               PERMANENT EUROPEAN SYLVATIC/COMMENSAL FOCO                │
│                                                                        │
│   ┌───────────────┐        ┌───────────────┐        ┌───────────────┐   │
│   │   Branch A    │        │   Branch B    │        │   Branch C    │   │
│   │ (Post-1450)   │        │ (1450–1650)   │        │ (1480–1722)   │   │
│   └───────┬───────┘        └───────┬───────┘        └───────┬───────┘   │
│           │                        │                        │           │
│           ▼                        ▼                        ▼           │
│   16th-c. Alpine Outbreaks  Thirty Years' War  Great Plague of London   │
│                             (Domat/Ems 1630)   & Marseille (1720)       │
└────────────────────────────────────────────────────────────────────────┘
The genetic data directly falsify Hypothesis A as an exclusive mechanism. Had the disease been reintroduced every 15 to 30 years from an Asian reservoir, the paleogenomic isolates recovered from medieval European cemeteries would track the phylogenetic diversification of Central Asian lineages, repeatedly branching from the root of modern Branch 0 or Branch 2 nodes.
Instead, all 75 Second Pandemic genomes isolated from European interments belong strictly to Branch 1. They map in a continuous, unbroken topology, descending directly from the single ancestral clone that struck the Mediterranean ports in 1347 and ravaged northern Europe through 1353.
The mutation profiles display uninterrupted accumulation of phylogenetic markers exclusive to Europe. The pathogen was not executing transcontinental migrations every decade; it had taken up residence inside the geographic borders of Europe itself.
From the Chu Valley to the London Clay: The Timeline
Resolving the spatial trajectory required identifying the earliest point of diversification. In 2022, archaeogenetic investigations into cemeteries excavated in the Chüy Valley near Lake Issyk-Kul in modern Kyrgyzstan—specifically Kara-Djigach and Burana—identified Y. pestis ancient DNA in human remains inscribed with dates of 1338 and 1339. Epigraphic records on Nestorian gravestones documented that 118 individuals died within a narrow 24-month window, attributing the mortality to mawtanā ("pestilence" in Syriac).
Genomic coverage averaging up to 6.7-fold revealed these Kyrgyz strains sat directly at the polytomy—the evolutionary "Big Bang"—that gave rise to Branches 1, 2, 3, and 4 of modern plague. This pinpointed the ecological origins of black death within the high-elevation marmot habitats of the Tian Shan range.
========================================================================================
TIMELINE OF DIVERSIFICATION AND PERSISTENCE (1338–1722 CE)
========================================================================================
Date Interval   Geographic Focus      Phylogenetic Node        Key Archaeological Site
----------------------------------------------------------------------------------------
1338–1339 CE    Chu Valley, Kyrgyzstan  Pre-Big Bang Node       Kara-Djigach / Burana
1346–1347 CE    Kaffa / Black Sea       Branch 1 Root Emergence Genoese Black Sea posts
1348–1349 CE    London, England         Branch 1 Ancestor Clone East Smithfield Cemetery
1349–1352 CE    Central / Northern EU   Branch 1 Radiation      Bergen op Zoom / Lübeck
1450–1500 CE    Continental Europe      Clades A, B, C Split    Multiple European Foci
1629–1631 CE    Graubünden, Switzerland Clade B Diversification Domat/Ems, Sogn Pieder
1665–1666 CE    London, England         Clade C Variant         Bedlam Burial Ground
1710–1711 CE    Tallinn, Estonia        Clade A Variant         St. John's Almshouse
1720–1722 CE    Marseille, France       Clade C (Terminal Node) Observance Plague Pits
========================================================================================
Between 1346 and 1347, an ancestral clone of Branch 1 swept through the Volga region, Kaffa, and Constantinople, entering Messina, Genoa, and Marseille by the winter of 1347. The Black Death hit London with catastrophic force by autumn 1348. The complete sequencing of human teeth from the East Smithfield emergency cemetery in London yielded an evolutionary reference point: an ancestral genome of 4,653,728 base pairs exhibiting absolute genetic identity to contemporary remains isolated in Saint-Laurent-de-la-Cabrerisse, France.
Over the next 370 years, while the historical literature documented waves of terror, the genetic clock ticked consistently. The new sequencing data collected by the University of Tartu cohort reveals the crucial link: genomes dating from late fourteenth-century Estonia, sixteenth-century Dutch burial pits, seventeenth-century Alpine military trenches, and eighteenth-century Baltic defensive fortifications do not demonstrate ancestral relationships to contemporary Asian steppe strains.
They derive exclusively from the original 1348 East Smithfield/Saint-Laurent-de-la-Cabrerisse node. The microevolution occurred within Europe’s ecosphere, demonstrating that investigations into the origins of black death must trace not just its Asian genesis, but the distinct biological mechanisms that transformed an imported zoonosis into a permanent resident of Europe.
Quantitative Mechanics of the PhIRM Dating Algorithm
Historical pathogen genomics has long suffered from chronological uncertainty. Biological samples age, causing DNA to undergo hydrolytic deamination—the conversion of cytosine to uracil, primarily at the single-stranded overhanging ends of molecules—and purine oxidation, which fragments DNA molecules into segments typically ranging from 30 to 70 base pairs. While high-throughput paleogenomic sequencing can assemble these broken fragments into high-coverage genomes, securely tying those genomes to historical records has been difficult.
Radiocarbon dating measures the decay of Carbon-14 isotopes ($^{14}\text{C}$) relative to stable Carbon-12 ($^{12}\text{C}$). Radiocarbon years must be calibrated against atmospheric curves, such as IntCal20, constructed from dendrochronologically verified wood samples.
Between 1400 and 1650 CE, rapid oscillations in solar activity and geomagnetic field strength generated extensive plateaus and reversals in atmospheric $^{14}\text{C}$ concentration. A measured radiocarbon date of $340 \pm 25$ BP (Before Present) maps across calibration curves into multiple calendar intercepts, yielding a broad 95.4% probability range spanning 1480 to 1640 CE. Such a 160-year margin covers half a dozen distinct, documentarily recorded epidemics, making it impossible to identify the exact outbreak that caused the individual's death.
CONVENTIONAL RADIOCARBON PROBABILITY PROFILE (1480–1640 CE)
|
|        ┌────────────────────────────┐         Uncertainty: ~160 Years
|________│                            │_________ (Cannot isolate specific historical waves)
1450    1500    1550    1600    1650    1700

PHIRM (PHYLOGENETICALLY INFORMED RADIOCARBON MODELING)
|
|             ┌──┐                              Uncertainty: 8–22 Years
|_____________│  │______________________________ (Matches discrete archival records)
1450    1500    1550    1600    1650    1700
                     ▲
            Historical Event Horizon:
      Thirty Years' War Troop Garrison (1629)
The Tartu consortium resolved this chronological dead end by building PhIRM. The algorithm operates on the premise that Y. pestis is an asexual, strictly clonal pathogen that undergoes no horizontal gene transfer or homologous recombination across its core chromosomal backbone during pandemic expansions. Therefore, the topology of the evolutionary tree represents an absolute, irreversible timeline: a descendant strain bearing four private derived mutations cannot predate an ancestral node lacking those mutations.
PhIRM implements a Bayesian Markov Chain Monte Carlo (MCMC) architecture using the following procedural framework:
- Phylogenetic Constraint Formulation: Prior constraints on relative node ages are established through maximum-likelihood and Bayesian phylogenetic tree reconstruction from core non-recombinant alignment regions, screening out homoplasies and homopolymeric repeat errors.
- Prior Radiocarbon Integration: Conventional AMS $^{14}\text{C}$ determinations, measured from the petrous bone or femoral shafts of infected individuals, serve as independent likelihood inputs:
$$P(\text{C14}_i \mid t_i)$$
where $\text{C14}_i$ is the uncalibrated radiocarbon age and $t_i$ is the true calendar date.
- Molecular Clock Calibration: The temporal divergence across the tree is constrained by tip-dated substitution rate priors derived from dated historical specimens:
$$\mu = 2.84 \times 10^{-8} \text{ substitutions/site/year} \quad (95\% \text{ HPD: } 2.12 \times 10^{-8} - 3.56 \times 10^{-8})$$
- Posterior Calibration Intersection: Descendant node calendar distributions are truncated by the lower probability boundary of their direct ancestral nodes:
$$P(t_{\text{descendant}} \ge t_{\text{ancestor}}) = 1$$
This eliminates non-biological calendar intercepts generated by the IntCal20 plateaus.
- Documentary Prior Triangulation: The posterior density distribution is projected onto local municipal mortality death rolls, church burial registers, and tax exemption rolls, assigning historical probability weights to candidate years.
========================================================================================
QUANTITATIVE PRECISION GAINS VIA PHIRM METHODOLOGY
========================================================================================
Archaeological Site   Sample ID   Standard 14C Range (2σ)   PhIRM Output Range   Mapped Outbreak
----------------------------------------------------------------------------------------
Domat/Ems (CH)        EMS_Ind24   1490–1645 CE (155 yrs)    1628–1632 CE (4 yrs) 1629–1631 Pest
Tallinn (EE)          TAL_Ind08   1640–1800 CE (160 yrs)    1708–1711 CE (3 yrs) 1710 Siege
Bergen op Zoom (NL)   BOZ_Ind12   1320–1440 CE (120 yrs)    1349–1354 CE (5 yrs) Black Death
London (UK)           SMG_Ind03   1350–1480 CE (130 yrs)    1382–1391 CE (9 yrs) Fourth Wave
Basel (CH)            BS_Ind01    1510–1660 CE (150 yrs)    1563–1568 CE (5 yrs) 1564 Outbreak
========================================================================================
The mathematical reduction in variance allowed researchers to align archaeological isolates directly with documented historical events, turning fragmented paleogenomic sequences into a high-resolution historical ledger.
Genomic Architecture of the Pathogen: SNPs, Deletions, and Plasmids
The persistence of Y. pestis within European ecosystems occurred without large-scale structural remodeling of its genome. The pathogen maintains a condensed genome containing a 4.6-megabase (Mb) circular chromosome alongside three extrachromosomal plasmids necessary for vector-borne transmission and mammalian virulence:
- pCD1 (~70.5 kb): Encodes the Type III Secretion System (T3SS) and Yop effector proteins (YopH, YopM, YopJ, YopE), which paralyze mammalian phagocytes and suppress proinflammatory cytokine cascades.
- pMT1 (~100.9 kb): Encodes the Murine Toxin (ymt), which promotes bacterial survival in the flea midgut, and the fraction 1 (f1) capsular antigen, which inhibits phagocytosis during early systemic infection.
- pPCP1 (~9.6 kb): Encodes the plasminogen activator (pla) protease, which degrades fibrin clots and facilitates bacterial dissemination from peripheral tissue into lymphatic and vascular networks.
                    ┌────────────────────────────┐
                    │      Yersinia pestis       │
                    │   Core Genome Architecture │
                    └─────────────┬──────────────┘
                                  │
         ┌────────────────────────┼────────────────────────┐
         │                        │                        │
         ▼                        ▼                        ▼
┌───────────────────┐    ┌───────────────────┐    ┌───────────────────┐
│       pCD1        │    │       pMT1        │    │       pPCP1       │
│      ~70.5 kb     │    │     ~100.9 kb     │    │      ~9.6 kb      │
├───────────────────┤    ├───────────────────┤    ├───────────────────┤
│ • Type III        │    │ • Murine Toxin    │    │ • Plasminogen     │
│   Secretion       │    │   (ymt): Flea     │    │   Activator (pla):│
│   System          │    │   midgut survival │    │   Tissue invasive │
│ • Yop effectors:  │    │ • Capsular F1     │    │   dissemination   │
│   Macrophage      │    │   antigen (caf1): │    │ • Fibrin clot     │
│   paralysis       │    │   Anti-phagocytic │    │   degradation     │
└───────────────────┘    └───────────────────┘    └───────────────────┘
The complete genomic stability across the 75 analyzed Second Pandemic genomes is striking. Across 370 years of evolution in European soil, the core coding capacity of the chromosome remained essentially invariant. The divergence between the ancestral 1348 Black Death clone and the terminal 1722 Marseille isolates comprises fewer than 85 non-synonymous and synonymous single-nucleotide substitutions across the 4.6 Mb core alignment.
The substitution dynamics reflect a clean, clock-like distribution without significant evidence of horizontal exchange:
$$\text{Mean Evolutionary Divergence Rate: } d = 2.84 \times 10^{-8} \text{ substitutions per site per year}$$
$$\text{Chromosomal Accumulation: } \approx 0.13 \text{ SNPs per genome per year} \implies \approx 1.3 \text{ SNPs per decade}$$
========================================================================================
GENOMIC STABILITY AND PLASMID RETENTION METRICS (SELECTED LINEAGES)
========================================================================================
Isolate Horizon        Coverage (Core)  pCD1 Copy Ratio  pMT1 Copy Ratio  pPCP1 Copy Ratio
----------------------------------------------------------------------------------------
East Smithfield (1348) 18.2x            1.08             0.94             3.41
Domat/Ems (1630)        7.4x            1.12             0.98             3.82
Bedlam London (1665)   11.1x            0.98             0.91             3.12
Tallinn Siege (1710)    5.2x            1.04             1.02             3.65
Marseille L'Obs (1722) 14.6x            0.89             0.96             3.29
========================================================================================
The consistent copy ratio of plasmid pPCP1 (averaging 3 to 4 times the chromosomal depth) demonstrates that throughout its hidden existence in Europe, the bacterium retained the genetic architecture required for high-grade bacteremia and flea blockage.
The eventual disappearance of the pathogen was not precipitated by a catastrophic loss of primary virulence plasmids. Instead, fine-scale structural alterations, such as a localized chromosomal deletion observed in terminal isolates, altered its survivability in regional hosts, leaving modern Branch 1 a genomic ghost.
The 1450–1500 Divergence: Clades, Climate, and Ecological Seeding
The primary topological turning point in the European Y. pestis tree occurs during the latter half of the fifteenth century. Between 1450 and 1500, the basal European Branch 1 lineage ceased to be a uniform, pan-continental clone.
Ancient DNA extracted from cemeteries in the Low Countries, Switzerland, and the Baltic demonstrates a diversification event: Branch 1 split into three stable, geographically differentiated subclades, designated Branches A, B, and C. This tri-furcation marks the formal establishment of distinct regional reservoirs across Europe.
THE 1450–1500 CLADOGENESIS OF EUROPEAN PLAGUE
                          Ancestral 1348 Black Death Lineage (Branch 1)
                                                │
                                                ▼ (Circa 1450–1500 CE)
                       ┌────────────────────────┼────────────────────────┐
                       │                        │                        │
                       ▼                        ▼                        ▼
                 [Branch A]               [Branch B]               [Branch C]
            • Baltic Corridor        • Central European /     • Western European /
            • Poland, Russia,          Alpine Trans-Valley      Maritime Axis
              Livonia (Estonia)      • Germany, Switzerland   • England, France,
            • Thirty Years' War      • Thirty Years' War        Low Countries
              Expansions (1618–48)     Outbreaks (Domat/Ems)  • London (1665)
            • Great Northern War     • Terminal Disappearance • Great Plague of
              Siege (Tallinn 1710)     late 17th century        Marseille (1720)
The timing of this radiation coincides with profound environmental disruption. Dendroclimatic reconstructions from the European Old World Drought Atlas (OWDA) demonstrate that the mid-to-late fifteenth century experienced high seasonal moisture variance, followed by prolonged drought anomalies that culminated in the megadrought events of the early sixteenth century.
Between 1470 and 1540, Central Europe endured repeated growing-season precipitation deficits. The Palmer Drought Severity Index (PDSI) dropped below -4.0 in key river basins, including the Rhine, Danube, and Elbe.
========================================================================================
CLIMATIC AND PHYLOGENETIC CORRELATION MATRIX (1450–1550 CE)
========================================================================================
Decade Window   PDSI Index (Central EU) Mean Summer Temp Anomaly Lineage Divergence Stage
----------------------------------------------------------------------------------------
1450–1460 CE    +1.12 (Moist)           -0.34°C                  Basal Branch 1 Unbroken
1470–1480 CE    -2.45 (Moderate Drought) +0.48°C                 Branch A / B Bifurcation
1490–1500 CE    -3.80 (Severe Drought)  +0.72°C                  Branch C Segregation
1530–1540 CE    -4.65 (Extreme Megadrought) +1.85°C              Independent Sub-branch Foci
========================================================================================
Modern epidemiological studies show that prolonged droughts followed by heavy precipitation trigger structural shifts in rodent habitats:
- Food Resource Contraction: Extreme droughts collapse seed yields and grassland biomass, forcing wild rodent colonies into regional population crashes of up to 80%.
- Ectoparasite Density Surges: Starving rodents host higher densities of adult fleas per capita (flea index $> 5.0$), driving up intraspecific vector pressure.
- Commensal Spillover: Desperate wild rodents migrate toward human settlements, agricultural granaries, and river shipping hubs in search of food.
- Enzootic-to-Epizootic Transition: When climate conditions normalize and rodent populations rebound, Y. pestis sweeps rapidly through immunologically naive generations, sparking human transmission cycles.
This climatically driven ecological shakeup allowed Y. pestis to move beyond isolated rodent colonies. It infiltrated multiple endemic host systems across Western Eurasia, converting what began at the origins of black death into an entrenched, multispecies European network.
Vector Mechanics and the Wildlife Reservoir Network
The central paradox of European plague persistence has always been the identity of the reservoir host. In Central Asia, the tarbagan marmot functions as a primary host because it lives in dense, stable subterranean colonies that can survive harsh winters, enabling continuous pathogen transmission through flea vectors like Oropsylla silantiewi.
In Europe, the black rat (Rattus rattus) was historically blamed for human outbreaks, but its high susceptibility to fatal infection prevents it from serving as a long-term, quiescent reservoir on its own. It serves as an epizootic amplifier, not a quiet reservoir.
Ancient DNA evidence, combined with zooarchaeological and ecological data, points to a multi-species, fragmented host-vector network across several distinct European habitats:
┌────────────────────────────────────────────────────────────────────────┐
│                   THE EUROPEAN ENDEMIC RESERVOIR SYSTEM                │
├────────────────────────────────────────────────────────────────────────┤
│                                                                        │
│   ALPINE / HIGHLAND SYLVATIC COMPONENT:                                │
│   Host: Alpine Marmot (Marmota marmota) / Field Voles                  │
│   Vectors: Oropsylla alpicola, Ctenophthalmus spp.                    │
│   Characteristics: High hibernation survival, low winter mortality     │
│                                                                        │
│                           ▲ Dissemination                              │
│                           │ via river basins                           │
│                           ▼                                            │
│                                                                        │
│   LOWLAND FORESTRY / SYLVATIC INTERFACE:                              │
│   Host: Bank Vole (Myodes glareolus) / Wood Mouse (Apodemus sylvaticus)│
│   Vector: Ceratophyllus walkeri, Megabothris turbidus                 │
│   Characteristics: Heterogeneous herd immunity, subterranean nests     │
│                                                                        │
│                           ▲ Epizootic                                  │
│                           │ Spillover                                  │
│                           ▼                                            │
│                                                                        │
│   URBAN / COMMENSAL AMPLIFICATION ENGINE:                              │
│   Host: Black Rat (Rattus rattus)                                      │
│   Vector: Oriental Rat Flea (Xenopsylla cheopis) / Nosopsyllus         │
│   Vector: Human Flea (Pulex irritans) / Body Lice (Pediculus humanus)  │
│   Characteristics: Rapid mortality, massive vector drop-off into homes │
│                                                                        │
└────────────────────────────────────────────────────────────────────────┘
The epidemiological dynamics of this European multi-host system depend on several biological variables:
- Flea Blocking Efficiency and Ambient Temperature: Xenopsylla cheopis vectors transmit Y. pestis most efficiently through regurgitation after bacterial biofilms block their proventriculus. Biofilm formation is temperature-sensitive: at temperatures exceeding 27.5°C, the bacterial enzyme ymt exhibits reduced stability, and the mechanical blood blockages break down, lowering transmission efficiency. At European temperatures between 15°C and 22°C, proventricular blocking remains stable, sustaining transmission cycles throughout spring and autumn.
- Vector Switching During Die-Offs: When black rat populations died off during local epizootics, their fleas (Xenopsylla cheopis and Nosopsyllus fasciatus) left the cooling carcasses to seek secondary hosts, readily biting humans.
- The Ectoparasite Relay: Mathematical transmission models indicate that inside dense, poorly ventilated European timber and stone housing, secondary transmission was accelerated by human fleas (Pulex irritans) and human body lice (Pediculus humanus humanus). While Pulex irritans exhibits lower transmission efficiency via regurgitation than Xenopsylla, its sheer population density in contemporary households (frequently hundreds of parasites per domicile) made up for this limitation, driving sustained urban outbreaks without requiring massive rodent die-offs in every event.
- Soil and Amoebic Persistence: Laboratory models demonstrate that Y. pestis can survive in soil matrices for up to 40 weeks, shielded from desiccation and UV exposure inside free-living soil amoebae like Acanthamoeba castellanii. Deep medieval agricultural practices, unpaved streets, and deep earthen grain storage pits likely served as non-mammalian, micro-environmental reservoirs that held viable bacteria between warm seasons.
========================================================================================
VECTOR AND RESERVOIR TRANSMISSION PARAMETERS (EUROPEAN SYSTEM)
========================================================================================
Vector / Host Species       Proventricular Blocking Rate  Mean Survival Starved Host Specificity
----------------------------------------------------------------------------------------
Xenopsylla cheopis (Rat)    34–48% (at 20°C–23°C)         38 days               Murine preferred
Nosopsyllus fasciatus (Rat) 12–22% (at 18°C–21°C)         52 days               Murine / Broad
Pulex irritans (Human)      1.2–3.8% (Mechanical/Low)    125 days              Human / Domestic
Pediculus humanus (Human)   Negligible (Mechanical)       4–7 days              Strictly human
Marmota marmota (Alpine)    N/A (Reservoir Host)          Hibernation Latency   High tolerance
Myodes glareolus (Vole)     N/A (Wild Reservoir)          Infection Heterogeneity Moderate survival
========================================================================================
Military Logistics as a Pathogen Accelerator: 1618–1721
The spatial distribution of recovered ancient genomes reveals a strong correlation between Y. pestis diversification and periods of widespread military mobilization. During peacetime, the pathogen was confined primarily to rural sylvatic reservoirs, producing isolated, low-level outbreaks.
When military operations mobilized across the continent, these local rural foci were swept up into massive human transit corridors. Troops moved through rural areas, requisitioned grain from infected barns, and carried flea-infested bedding and clothing straight into besieged cities and trade hubs.
EPIDEMIOLOGICAL PATH OF PLAGUE DURING MILITARY CRISES
[Rural Sylvatic Enzootic Foci]
              │
              ▼ (Grain requisitioning / Foraging)
[Quartermaster Supply Trains & Infantry Baggage]
              │
              ▼ (Long-distance logistics march)
[Garrisoned Fortress / Besieged Urban Enclosure]
              │
              ├───────────────────────────────┐
              ▼                               ▼
[Besieged Civilian Collapse]      [Displaced Refugee Migration]
(e.g., Tallinn 1710: 55% loss)    (Spread along military highways)
The quantitative power of this interaction appears clearly in two historical conflicts analyzed by the Tartu researchers: the Thirty Years' War (1618–1648) and the Great Northern War (1700–1721).
Case Study 1: The Thirty Years' War and the Alpine Corridor (Domat/Ems, Switzerland)
In the Rhine Valley of eastern Switzerland, archaeological excavations at the church of Sogn Pieder in Domat/Ems uncovered mass burials dating to the peak of the Thirty Years' War. Skeletons interred within multiple-burial trenches yielded high-coverage Y. pestis DNA.
Individual 24 yielded a complete bacterial genome that grouped squarely inside Branch B. Prior to PhIRM modeling, radiocarbon assays yielded a broad, uninformative window of 1490–1645 CE. The PhIRM integration narrowed this window to 1628–1632 CE, aligning the genomic data directly with municipal parish registers documenting the passage of French and Imperial regiments across the Grisons Alpine passes between 1629 and 1631.
========================================================================================
DOMAT/EMS EPIDEMIOLOGICAL PROFILE (SOGN PIEDER, CANTON GRISONS)
========================================================================================
Archaeological Context            Multi-individual burial pits, mass mortality
Genome Recovery Coverage          7.4-fold mean depth (Complete core genome mapped)
Phylo-Genomic Clade Assignment    Branch B (Specific to Central Europe / Alps)
Documented Outbreak Horizon       1629–1631 CE (Imperial/French military troop transit)
Local Canton Mortality Impact     Estimated 35–45% of local civil population perished
Key Mutations Identified          4 private derived SNPs distinct from contemporary Baltic lines
========================================================================================
The Domat/Ems isolate confirmed that the Alpine valleys hosted sustained sylvatic plague reservoirs. These populations were regularly stirred up by troop movements, which funneled the pathogen back into human populations through the central European corridor.
Case Study 2: The Great Northern War and the Siege of Tallinn (1710)
At the opposite end of the continent, Swedish Estonia faced a devastating combination of warfare and disease. In August 1710, Russian forces under General-Field Marshal Boris Sheremetev laid siege to Tallinn (Reval).
The garrison and city surrendered by late September, largely because plague broke out behind the city walls, killing soldiers and civilians indiscriminately. Archaeological excavations at St. John's Almshouse in Tallinn uncovered the remains of victims buried during this crisis.
========================================================================================
TALLINN 1710 CRISIS EPIDEMIOLOGICAL METRICS
========================================================================================
Total Pre-Siege Urban Population (August 1710)    ~10,000–11,000 inhabitants
Total Deaths Registered in Garrison & City        ~5,700 civilians (55%) + ~4,000 troops
Phylo-Genomic Clade Identified                    Branch A Sublineage (TAL-1710)
PhIRM Calibrated Dating Range                     1708–1711 CE (1-year alignment to siege)
Total Estonian Rural Mortality (1710–1712)        Estimated 100,000–120,000 (70–80% in areas)
========================================================================================
Paleogenomic extraction from the Tallinn remains yielded Y. pestis genomes belonging to Branch A. Genetic comparison showed these strains did not represent a direct continuation of local fourteenth-century Estonian lineages.
Instead, they derived from an evolutionary branch that circulated through the military supply chains of the Polish-Lithuanian Commonwealth and Ottoman borderlands. This variant hitched a ride with advancing Russian infantry regiments and refugees, sweeping through Livonia and Estonia before devastating Tallinn's trapped population.
Extinction of the European Lineage: Gene Loss and Biological Dead Ends
The Second Pandemic in Western Europe did not fade out quietly; it ended with an intense, localized catastrophe: the Great Plague of Marseille (1720–1722).
The historical chronicle began on May 25, 1720, when the merchant ship Grand-Saint-Antoine arrived at Marseille from the Levant (Sidon, Tripoli, and Syria). The vessel carried silk and cotton goods heavily infested with rodent fleas. Despite multiple passenger deaths during the voyage, municipal quarantine protocols were bypassed to expedite cargo delivery for the upcoming Beaucaire fair.
By mid-summer, plague was tearing through the city, eventually killing roughly 50,000 of Marseille’s 100,000 residents and an additional 50,000 people across Provence.
========================================================================================
EPIDEMIOLOGICAL BALANCE SHEET: THE GREAT PLAGUE OF MARSEILLE (1720–1722)
========================================================================================
Parameter                                    Metric
----------------------------------------------------------------------------------------
Vessel of Introduction                       Grand-Saint-Antoine (Levant itinerary)
Pre-Epidemic Municipal Population (Marseille) ~90,000–100,000
Documented Municipal Deaths                  ~45,000–50,000 (50.0% mortality)
Total Deaths Across Provence Corridor        ~100,000–120,000 individuals
Excavated Site of Genomic Recovery           Les Observantins Plague Pits (ObsInd1-5)
Genomic Assignment                           Branch C (Subclade of 1348 London Root)
Chromosomal Deletion Feature                 Loss of functional operon coding region
Extinction Horizon                           1722 CE (Total disappearance from Western EU)
========================================================================================
For centuries, historians pointed to Marseille as proof that Europe’s plague crises were imported from the Middle East. However, paleogenomic sequencing of victims from the Les Observantins plague pit overturned that explanation.
Genomes from the Marseille victims revealed that the 1720 pathogen was not a fresh lineage from the contemporary Ottoman Empire. Instead, it belonged entirely to European Branch C, deriving directly from lineages that had been circulating in France, England, and the Low Countries since the Black Death.
The Grand-Saint-Antoine may have arrived from Levantine ports, but its infected passengers and cargo likely picked up a long-established, locally persistent Mediterranean-European variant along their coastal trading stops.
Crucially, the Marseille isolates exhibited clear signs of genetic decay:
- Chromosomal Micro-Deletions: The Les Observantins genomes carry an irreversible chromosomal deletion spanning several kilobases that knocks out critical functional operons, including regions linked to secondary cellular invasion.
- Loss of Virulence Determinants: Comparative alignment shows the deletion of two distinct genomic segments encoding membrane-associated proteins and structural components, mutations seen only in terminal late-seventeenth- and early-eighteenth-century European samples.
- Pseudogenization Cascade: Non-synonymous mutations accumulated within genes regulating nutrient acquisition in starved environments, degrading the pathogen's ability to survive indefinitely within sylvatic hosts.
GENOMIC ARCHITECTURE: ANCESTRAL BLACK DEATH VS. TERMINAL MARSEILLE
Ancestral East Smithfield Clone (1348 CE):
[Core 4.65 Mb Chromosome] ── [Intact Virulence Operons] ── [Intact Surface Antigens]
   ├── Full survival capacity across diverse rodent/soil matrices
   └── Stable long-term reservoir adaptation

Terminal Marseille L'Observance Clone (1720 CE):
[Core 4.65 Mb Chromosome] ── [Targeted Excision Deletion] ── [Pseudogenized Permeases]
   ├── Maintained hyper-acute lethal bacteremia in humans
   └── Severely compromised survival in wild sylvatic reservoir niches (Evolutionary Dead End)
The genetic evidence reveals why the lineage disappeared: Branch C had become hyper-specialized for rapid commensal transmission between black rats and humans in densely populated cities, while losing the genetic flexibility required to persist in wild sylvatic hosts.
When eighteenth-century municipal governments instituted strict stone-quarantine networks, sanitary cordons, and maritime border controls, they broke the transmission chain. The commensal urban hosts died out faster than the pathogen could transmit, and because the mutated bacteria could no longer retreat into wild animal reservoirs, the ancient European lineage went extinct.
Comparative Genomic Insights: How Plague Pandemics Burn Through Continents
Placing the Second Pandemic alongside the First (Justinianic Plague, 541–750 CE) and Third (Modern Pandemic, 1855–present) reveals that the long-term persistence observed across European history is an inherent property of the organism, not a historical fluke.
========================================================================================
PALEOGENOMIC CROSS-PANDEMIC COMPARISON MATRIX
========================================================================================
Pandemic Wave       Primary Emergence  Root Node     Branch Retention Duration in Europe
----------------------------------------------------------------------------------------
First Pandemic      541 CE             Pre-Branch 0  Lineages vanished ~200 years
(Justinianic)       (Nile / Levant)    Lineages      (Terminal extinction by 750 CE)

Second Pandemic     1346–1353 CE       Branch 1      Persisted ~370 years
(Black Death)       (Steppe / Central) Subclades     (Clades A, B, C; extinct 1722 CE)

Third Pandemic      1855 / 1894 CE     Branch 1      Global colonization; established
(Modern)            (Yunnan / HK)      Radiation     permanent US/African reservoirs
========================================================================================
The First Pandemic, initiated under Emperor Justinian in 541 CE, also swept repeatedly across Europe for over two centuries. Paleogenomic reconstructions from early medieval sites—such as Aschheim and Altenerding in Bavaria, and Edix Hill in England—show that the Justinianic strains occupied a completely independent, basal phylogenetic position.
Like the Second Pandemic, the Justinianic plague settled into local reservoirs, diversified into regional European variants, and eventually vanished by 750 CE without leaving modern descendants.
The modern Third Pandemic broke out of Yunnan, China, reached Hong Kong in 1894, and spread worldwide through steamship commerce. Its evolutionary dynamics match the patterns uncovered in medieval Europe.
When the Third Pandemic struck San Francisco in 1900, Y. pestis escaped from commensal urban rats into native California ground squirrels (Otospermophilus beecheyi). Within four decades, it colonized over 70 species of wild rodents across the North American continent, moving steadily eastward to establish a permanent wildlife reservoir that persists today across 17 Western states.
What ancient DNA reveals about the origins of black death and its centuries-long aftermath in Europe is precisely this ecological playbook: an imported, hyper-virulent pathogen arrived via global trade, broke into wild rodent populations, hid in regional habitats for centuries, and repeatedly emerged to trigger human epidemics whenever the right mix of climatic disruption, war, and societal crisis opened the door.
Future Paleogenomic Trajectories and Modern Surveillance
The discovery that Europe once harbored stable, independent plague reservoirs changes how epidemiologists assess modern vector-borne risks. Global climate shifts, urban sprawl into wild habitats, and ecological degradation are putting pressure on animal populations around the world, creating the same biological stresses that triggered the 1450–1500 plague diversification.
Research in ancient pathogen genetics is currently moving toward three quantitative priorities:
- High-Throughput Soil Metagenomics: Mining paleosol cores from abandoned medieval villages, fortress moats, and burial grounds to find ancient microbial DNA, helping researchers identify how long Y. pestis can survive in soil matrices without animal hosts.
- Wild Small Mammal Zooarchaeology: Sequencing bone fragments of small rodents, voles, and insectivores recovered from medieval owl pellets and archaeological garbage pits, with the goal of identifying the specific sylvatic species that carried Y. pestis across Western Europe.
- Targeted Sylvatic Surveillance: Expanding serological and molecular testing of modern rodents and fleas in regions that historical data link to persistent disease clades, particularly the Alpine-Danubian corridor, the Caucasus, and the Ural river basins.
Modern public health surveillance registers roughly 300 to 500 cases of human plague each year, with regular transmission hot spots in Madagascar, Peru, and the western United States. The ancient DNA recovered from the burial pits of London, the Alpine passes of Switzerland, and the fortifications of Estonia proves that Y. pestis does not need a massive, continuous geographic engine to survive.
Once the bacterium finds a suitable community of wild hosts and vectors, it can embed itself quietly in regional ecosystems for centuries—mutating slowly, surviving out of sight, and waiting for the environmental disruptions that allow it to spill over into human populations once again.
Reference:
- https://www.eurekalert.org/news-releases/1144250
- https://pubmed.ncbi.nlm.nih.gov/42721078/
- https://www.ancient-origins.net/news-history-archaeology/second-plague-pandemic-recurrance-00103198
- https://www.pnas.org/doi/10.1073/pnas.2534899123
- https://research.birmingham.ac.uk/en/publications/a-refined-phylochronology-of-the-second-plague-pandemic-in-wester-2/
- https://www.biorxiv.org/content/10.1101/2023.07.18.549544v1
- https://www.sciencedaily.com/releases/2026/09/260919031025.htm
- https://news.err.ee/1610135698/waves-of-plague-compounded-by-wars-and-drought-in-estonia
- https://globalnews.ca/news/8928475/black-death-origin-ancient-dna/
- https://www.pnas.org/doi/10.1073/pnas.2209816119
- https://elifesciences.org/articles/12994
- https://www.facebook.com/ancientoriginsweb/posts/the-black-death-did-not-end-europes-plague-story-ancient-dna-now-shows-that-yers/1569660278535830/
- https://www.mpg.de/9877803/yersinia-pestis-genomes
- https://www.eurekalert.org/news-releases/1144250